Dietary supplement production turns an agreed formula into a traceable batch through controlled materials, processing, packaging and a documented release decision.
A formula describes what a product should contain. A production system must also explain which materials were used, how they were handled, what changed during processing and why the finished goods can move out of quarantine. Those are different questions. A convincing sample can establish a starting point, but it cannot describe every later manufacturing run. Understanding the handoffs between these questions makes a production plan easier to review before artwork and delivery dates become fixed.
The Formula Becomes An Instruction That Can Be Repeated
Separate The Product Concept From The Controlled Version
A development brief may describe a flavour, dosage form and intended market. Manufacturing needs a version-controlled definition that identifies ingredients, their required characteristics, quantities, processing steps and the intended package. The master manufacturing record connects these choices. It should be clear which revision is authorised and what happens to an older version when the formulation changes. An email approving a taste sample does not automatically approve a changed ingredient source, a different capsule shell or a new serving statement.
For example, moving a powder from a jar to a single-serve sachet changes more than the artwork. The fill operation, seal checks, coding space and handling conditions may need review. The OEM and ODM workflow provides a useful starting point for discussing that handoff, but the actual project still needs its own approved instructions. A catalogue example illustrates a possible format; it does not establish the final specification of a separate order.
Choose Checks That Match The Dosage Form
Capsules, tablets and powders do not share every manufacturing risk. A capsule project may need attention to fill behaviour and shell suitability. A powder project may need evidence about blend uniformity, flow and filling consistency. A tablet project adds compression behaviour and handling strength. The correct controls depend on the formula and process. Treating every format as the same sequence can leave a gap between a laboratory sample and commercial equipment.
Materials Need Both An Identity And A Disposition
A Supplier Document Is One Part Of The Receiving Decision
Receiving a material is not the same as releasing it for use. The receiving record should connect the delivered container to the purchase requirement, supplier, lot identity and inspection status. A certificate of analysis can support that decision, but a document is useful only when it belongs to the received material and addresses the relevant specification. The verification plan should explain what is checked directly and what supplier evidence is accepted under an established qualification process.
Material traceability also needs a practical physical system. If two bags look alike but have different lot numbers or approval states, the operator should not have to infer which one is available. Storage locations, status identification and the production issue record need to agree. Quarantined or rejected materials should not become available merely because a planning screen shows stock on hand.
Preserve The Connection When Containers Are Split
Opening a bulk container, weighing a portion and returning the remainder creates another identity handoff. The weighed portion must remain connected to its source lot, and the remainder needs an appropriate status and storage condition. When several input lots feed one finished batch, the record should preserve that relationship. A neat warehouse photograph cannot prove that the relationship exists; it is the movement history and controlled identification that make a later investigation possible.
A review of the manufacturing facility context can identify which areas deserve a closer discussion. The useful follow-up is a walkthrough of one material movement: receipt, sampling, approval, weighing and issue. That reveals whether responsibilities remain clear when work passes between warehouse and production staff.
Processing Records Explain What Actually Happened
The Planned Sequence And The Executed Sequence Must Connect
The master manufacturing record describes the authorised method. A batch-specific record captures the execution, including material lots, quantities, equipment identification, relevant processing observations and required checks. These records serve different purposes. A copied standard instruction with no execution information cannot explain a finished batch, while an operator log without an approved reference leaves uncertainty about what the process was supposed to achieve.
Dietary supplement production should make unusual events visible. A pause, an adjustment or a material discrepancy may be acceptable after assessment, but it should not disappear into a final signature. A meaningful record identifies the event, its potential effect, the action taken and the disposition. The objective is an understandable manufacturing history, not a record that appears perfect because exceptions were never recorded.
In-Process Results Need A Decision Attached
A measurement becomes useful when its acceptance rule and response are defined. A fill observation, compression check or blend sample should connect to a planned criterion and an action when the result is outside that criterion. Recording numbers without a decision can create a false sense of control. Equally, a generic statement that a line was inspected does not identify which characteristic was examined or what evidence supported continuation.
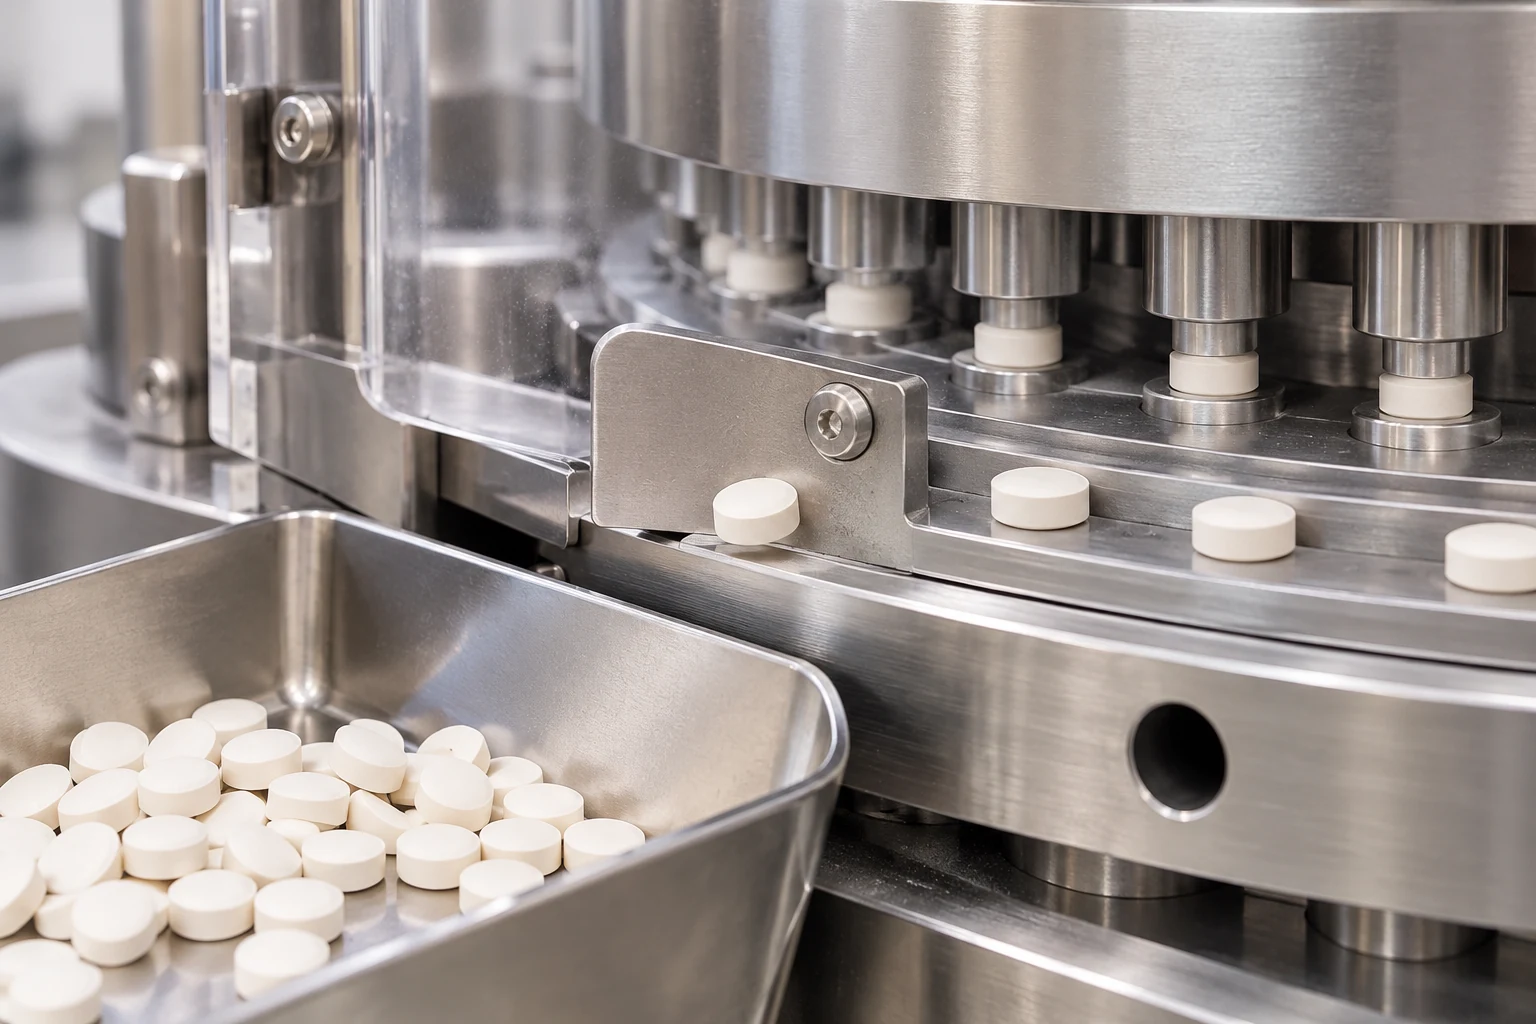
Reconciliation helps close the physical side of the process. The quantities issued, used, returned, rejected and packed should tell a coherent story within the approved procedure. An unexplained discrepancy deserves investigation even when the finished packages look correct. It may indicate a recording problem, material loss or an identity issue; the explanation must come from evidence rather than an assumed normal allowance.
Testing Answers Specific Questions About The Batch
A Result Has Meaning Only In Its Sampling Context
A laboratory result is tied to a sample, method and acceptance requirement. Before relying on it, establish what was sampled, when it was taken, how it represents the material or finished batch, and which question the method addresses. A result from an earlier development sample does not automatically represent the next commercial lot. Likewise, a test that confirms one characteristic does not prove every other quality attribute.
Batch release therefore requires more than attaching a laboratory file to a shipment. The reviewer needs to know whether required results are complete, whether the methods and specifications are appropriate, and whether unresolved events could affect the interpretation. When an external laboratory is involved, the sample identification and requested analysis must remain consistent with the project record.
For an Australian market project, first establish whether the intended presentation falls within food or therapeutic-goods requirements. TGA guidance distinguishes categories of complementary medicines, with different regulatory pathways. That classification cannot be determined from a generic supplement name alone. Ingredients, claims and presentation matter. A manufacturing discussion should identify the applicable pathway before assuming that an overseas label or certificate is sufficient.
The FDA dietary supplement manufacturing guidance provides a separate example of how specifications, production records and quality control responsibilities fit together in the United States. It is useful context for understanding the record chain, not a substitute for the requirements applicable to an Australian product. No catalogue listing, sample image or general manufacturing description establishes market authorisation.
Packaging And Release Close The Evidence Chain
Packaging preparation adds its own controlled identities: the approved artwork revision, container and closure, batch code and any required date information. A correct formula in the wrong labelled container is still a serious mismatch. Line clearance, component verification and reconciliation of printed materials should make it possible to explain which version was used and how unused or obsolete components were handled.
| Handoff | Question The Record Must Answer | Unresolved Gap To Investigate |
|---|---|---|
| Formula to manufacturing | Which authorised version governed this run? | Sample approval exists, but the production revision is unclear. |
| Warehouse to weighing | Which released input lots were issued? | Container identity or remaining-stock status is missing. |
| Processing to quality review | Were checks completed and exceptions resolved? | Results are present without criteria or disposition. |
| Bulk product to packaging | Which artwork and components were used? | Label approval does not match the packed version. |
| Quarantine to dispatch | Who authorised release on what evidence? | A shipping instruction is being treated as quality approval. |
A practical batch release review connects those handoffs instead of treating each file as an independent achievement. It establishes whether the required evidence is complete, whether any deviation remains unresolved and whether the finished goods have the correct identity. The release authority and escalation route should be agreed before a delivery deadline creates pressure to substitute a logistics decision for a quality decision.
For a new project, the development partnership overview can help frame responsibilities, while the available dosage formats clarify which production route needs discussion. Neither replaces the project record. A useful explanation follows one example lot from its inputs to its final disposition and states clearly where evidence is still pending.
Frequently Asked Questions
Is An Approved Sample Enough To Start A Commercial Run?
A sample can confirm selected development characteristics, but the production version, materials, processing instructions and packaging requirements still need approval. Any differences between sample preparation and commercial manufacture should be identified and assessed before the run begins.
Does A Certificate Of Analysis Replace Production Records?
No. It reports specified analytical information for an identified sample or lot. It does not by itself describe all materials, processing events, packaging controls or review decisions. Those records complement each other rather than perform the same function.
Why Can The Same Formula Need A Different Process?
Equipment, scale, material characteristics and packaging can change how a formula behaves. The authorised process should address the actual manufacturing conditions. A successful small preparation is a useful observation, but its performance at another scale needs appropriate evidence.
What Makes A Production Explanation Useful Before An Order?
It should identify the responsible parties, approved versions, important controls, expected records and the route for handling exceptions. A concrete example of material traceability is more informative than a general promise that every stage is monitored.
When Is A Finished Batch Ready To Ship?
When the required quality review and release decision have been completed under the applicable procedure, and dispatch requirements are also satisfied. Completion of packing or booking transport does not independently establish that release has occurred.
Dietary supplement production is best understood as a connected record of controlled decisions. Following one formula through materials, execution, testing and final disposition reveals what a production description can substantiate and which questions still need an answer.